
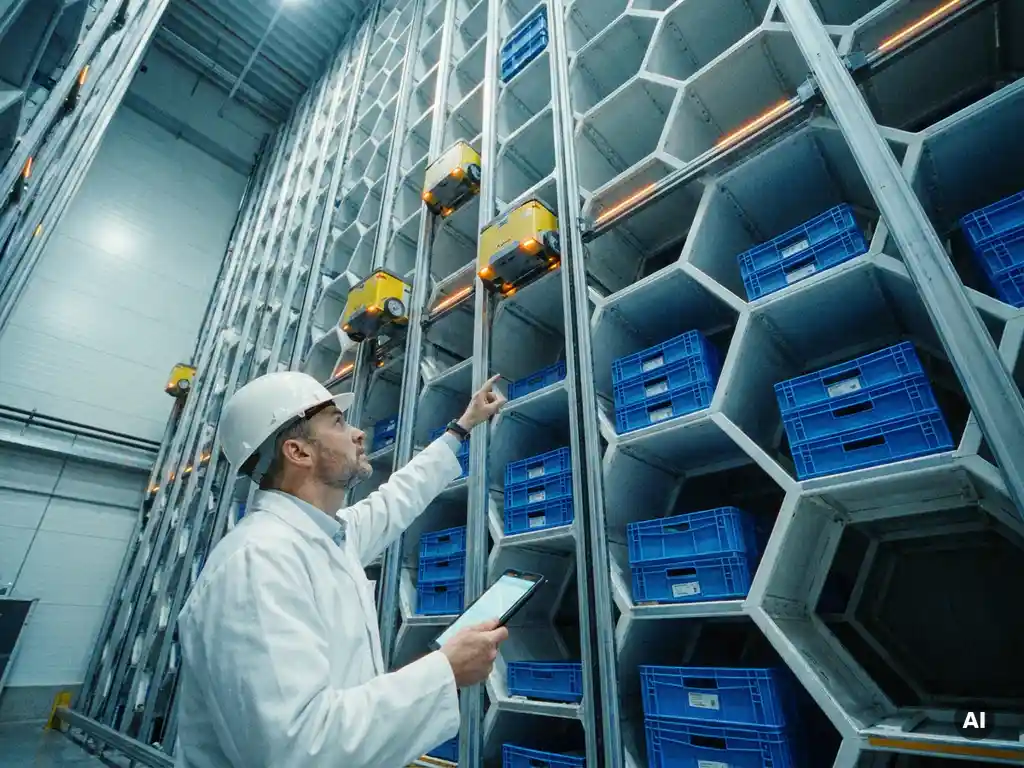
In 2026, regulatory agencies inspect pharmaceutical warehouses through a combination of announced and unannounced audits conducted by bodies such as the FDA, EMA member authority inspectors, and national competent authorities. Inspectors assess whether storage conditions, inventory controls, documentation practices, and distribution processes meet Good Distribution Practice (GDP) and Good Manufacturing Practice (GMP) standards. The sections below address the most common questions pharmaceutical warehouse operators face when preparing for or responding to a regulatory inspection.
What triggers a regulatory inspection of a pharmaceutical warehouse?
A pharmaceutical warehouse inspection can be triggered by a routine surveillance schedule, a product complaint, a reported deviation, a license application, or intelligence suggesting a compliance risk. Regulatory agencies operate both planned inspection programs and risk-based reactive programs, meaning a facility can receive an unannounced visit at any time if a credible concern exists.
Routine inspections are scheduled based on a facility’s risk classification, which considers factors such as the type of products handled, the volume of distribution, previous inspection history, and the complexity of operations. Facilities handling controlled substances, cold-chain products, or high-value biologics are typically inspected on shorter cycles than standard ambient-storage operations.
Reactive inspections are initiated when a regulator receives a complaint about product quality, a report of a falsified medicine in the supply chain, a serious adverse event with a suspected distribution link, or a whistleblower disclosure. Import alerts, border detentions, and international intelligence sharing between agencies such as the FDA and EMA member authorities can also prompt an unscheduled visit. License renewals and new facility approvals require a pre-approval inspection as a standard step.
What do inspectors actually examine during a pharmaceutical warehouse audit?
During a pharmaceutical warehouse audit, inspectors examine temperature and humidity monitoring records, inventory control systems, receiving and dispatch procedures, pest control logs, personnel training records, and the overall physical condition of the storage environment. The audit is designed to verify that every process that could affect product quality or supply chain integrity is controlled and documented.
Inspectors typically walk the facility before reviewing paperwork, looking for visible signs of poor storage conditions, damaged product, unlabeled materials, or unsecured controlled substances. They then move into records review, cross-checking whether documented procedures match observed practice. Discrepancies between written SOPs and actual operations are among the most common findings.
Physical storage conditions
Temperature mapping studies, calibration records for monitoring equipment, and alarm response logs are all subject to scrutiny. Inspectors verify that cold-chain products have remained within specification throughout their time in the facility and that any excursion was identified, investigated, and resolved with appropriate documentation.
Inventory and traceability controls
Batch number tracking, expiry date management, and the ability to rapidly locate or quarantine a specific lot are central to the audit. Inspectors may request a demonstration of how the facility would execute a product recall or field safety corrective action, testing whether inventory systems can isolate affected stock quickly and accurately.
How have GDP and GMP inspection standards changed for 2026?
In 2026, GDP and GMP inspection standards have evolved to place greater emphasis on data integrity, digital audit trail requirements, and supply chain transparency. Regulators increasingly expect facilities to demonstrate that electronic records are protected against unauthorized alteration, that system access is controlled and logged, and that data generated by automated systems is as auditable as paper-based records were previously.
The Falsified Medicines Directive (FMD) serialization requirements, already embedded in European distribution practice, are now being enforced with greater rigor, and inspectors are more likely to test the practical functionality of verification systems rather than simply confirming their presence. In the United States, the FDA’s Drug Supply Chain Security Act (DSCSA) traceability requirements have reached full enforcement maturity, meaning wholesale distributors and third-party logistics providers face detailed scrutiny of their electronic product tracing systems.
Sustainability and energy efficiency are not yet formal inspection criteria, but regulators are increasingly interested in whether environmental monitoring systems are robust enough to detect failures in temperature-controlled zones, particularly as climate-related risks to storage conditions receive more attention. The overall direction of travel is toward outcome-based inspection, where inspectors assess whether a facility can demonstrate control rather than simply produce a folder of signed documents.
What are the most common warehouse inspection findings that lead to citations?
The most common pharmaceutical warehouse inspection findings that result in citations include inadequate temperature excursion investigation, incomplete batch traceability records, insufficient personnel training documentation, failure to segregate quarantined or rejected stock from releasable inventory, and SOPs that do not reflect actual practice. These findings appear repeatedly across FDA warning letters and EMA inspection reports.
Temperature and environmental control failures are consistently among the top citations because the consequences for product quality are direct and serious. Facilities that cannot produce complete, continuous monitoring data for a specific storage period, or that cannot demonstrate a documented response to an alarm, face significant regulatory risk regardless of how well other systems perform.
Documentation gaps are the second major category. Inspectors look for evidence that every transaction, every deviation, and every corrective action has been recorded contemporaneously. Retrospective documentation, missing signatures, or records that cannot be linked back to a specific batch or order are all citation triggers. Personnel training records that are incomplete or that show operators performing tasks they have not been formally qualified to perform are a related and frequent finding.
Segregation failures, where quarantined, returned, or expired product is not physically or systematically separated from releasable stock, represent a direct product safety risk and are treated seriously. Facilities relying on manual location management are more vulnerable to this finding than those using software-directed inventory systems that enforce status-based access controls.
How do automated storage systems affect regulatory compliance assessments?
Automated storage systems can strengthen a pharmaceutical warehouse’s regulatory compliance position by providing software-directed inventory control, digitally logged retrieval events, and systematic enforcement of batch and expiry management rules. However, inspectors assess the complete implementation, not the hardware alone, meaning software validation, access controls, and audit trail integrity are all subject to scrutiny.
A well-implemented automated storage and retrieval system reduces the reliance on manual location management, which is one of the primary sources of segregation failures and traceability gaps. When a system directs every retrieval to a specific tote and logs the transaction against a batch number and timestamp, the audit trail is generated as a natural byproduct of normal operations rather than as a separate manual recording step.
Direct-access architectures are particularly relevant to pharmaceutical compliance because they eliminate the need to move or disturb surrounding inventory to reach a specific lot. In a system where every storage location is independently accessible, the risk of accidental cross-contamination, mispick, or unrecorded stock movement is structurally reduced. This is the type of operational control that inspectors look for when assessing whether a facility’s physical setup supports its documented procedures.
Regulators will examine whether the automated system has been formally validated, whether changes to software are controlled through a change management process, and whether the system generates complete, unalterable records. Facilities should be prepared to demonstrate that electronic records produced by the system meet the same integrity standards as paper records, and that access to the system is restricted and logged by individual user.
What documentation must a pharmaceutical warehouse produce during an inspection?
During a pharmaceutical warehouse inspection, a facility must be able to produce its distribution authorization or wholesale dealer license, standard operating procedures for all key processes, temperature monitoring records, batch traceability records, personnel training files, deviation and corrective action logs, supplier qualification records, and recall or product retrieval procedure documentation. The ability to locate and present these records quickly is itself an indicator of organizational control.
Inspectors typically request documentation in a specific sequence. They begin with the quality management system overview, including the site master file or equivalent facility description, then move to process-specific records for areas of interest identified during the facility walk. The speed and completeness with which a facility responds to document requests signals how well its quality system functions in practice.
Batch records and traceability documentation must demonstrate an unbroken chain from receipt to dispatch for any product an inspector selects. This includes goods-in records, storage location history, any intermediate handling or repackaging, and the dispatch record showing where the product was sent. For serialized products subject to FMD or DSCSA requirements, the electronic verification records form part of this chain and must be accessible in a format the inspector can review.
Deviation records and corrective action documentation are examined to assess whether the quality system is functioning as a genuine improvement mechanism or simply as a paper exercise. Inspectors look for evidence that deviations were investigated to root cause, that corrective actions were implemented within committed timelines, and that the effectiveness of those actions was subsequently verified.
How Hexxabotics supports pharmaceutical warehouse compliance
Hexxabotics provides a high-density automated storage and retrieval system designed to address the inventory control, traceability, and operational reliability requirements that pharmaceutical warehouse inspections assess most closely. For pharmaceutical distributors and fulfillment operations facing growing SKU counts, space constraints, and stricter regulatory scrutiny, the system offers a structurally compliant foundation for storage operations.
- Direct-access tote storage: Every storage location is independently accessible, eliminating the need to move surrounding inventory to retrieve a specific batch or lot, which reduces the risk of unrecorded stock disturbance and segregation failures.
- Digitally logged inventory movements: Each retrieval and storage event is recorded by the Hexxabotics Control System, supporting the audit trail requirements that inspectors examine during GDP and GMP assessments.
- Batch and expiry management support: Software-directed retrieval can support batch-controlled and expiry-aware workflows, helping facilities demonstrate systematic control over lot selection and FEFO compliance.
- Scalable capacity and throughput: Storage capacity and robot throughput can be scaled independently, allowing operations to grow without redesigning infrastructure or disrupting validated processes.
- Integration with WMS and orchestration layers: Standard API connectivity allows the system to operate within an existing warehouse management or execution system, preserving the data flows that compliance documentation depends on.
If your pharmaceutical warehouse operation is preparing for an inspection or evaluating how automation can strengthen your compliance position, contact Hexxabotics to discuss how the system can be configured for your specific regulatory and operational requirements.
Related Articles
- What is quarantine storage and when is it required in pharma warehouses?
- What is a GMP-compliant warehouse and how is it certified?
- What is direct-access storage and how does it support drug recalls?
- What documentation is required for controlled substance warehousing?
- What is the difference between CAPEX and OPEX in warehouse automation?